Our research
Chemical Dynamics: therory, computation and application

Research in the Geva group focuses on developing theoretical and computational approaches for simulating the dynamics of inherently quantum-mechanical dynamical processes underlying the performance and function of complex molecular systems. To this end, we combine a wide array of theoretical and computational strategies from quantum nonequilibrium statistical mechanics, the theory of quantum open systems, quantum master equations, the theory of stochastic processes, optical response theory, the path integral formulation of quantum mechanics, and semiclassical and mixed quantum-classical methods for simulating chemical dynamics.
Current research directions include:
Derivation of quantum master equations for systems coupled to multiple heat baths and driven by time-dependent fields, with emphasis on the effect of driving fields on the rate of energy transfer and decoherence and its relationship to thermodynamical consistency.
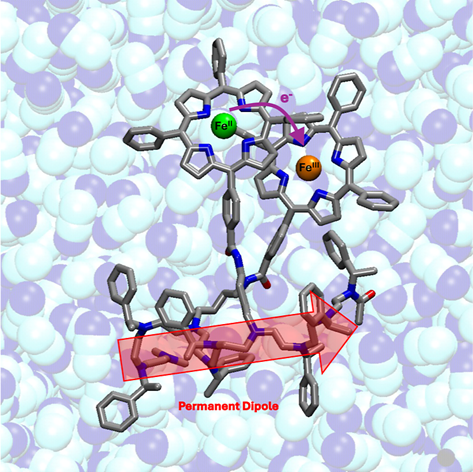
Simulation of charge transfer rates in emerging peptoid-based materials towards gaining a better understanding of their dependence on molecular and electronic structure and improving material performance.
Advancing the use of ion-trap devices as analog simulators of vibronic dynamics in molecular systems.
Using machine learning to address conceptual challenges and computational bottlenecks encountered in the simulation of chemical dynamics in complex molecular systems.
Exploring the interplay between chirality and the rate of decoherence.